AI | Bioinformatics | Deep Learning | News | Paper Summary | Structural Biology | Tools | Trending
Unlocking Molecular Dynamics: Dynaformer, a Graph-Based Deep Learning Model for Binding Affinity Prediction
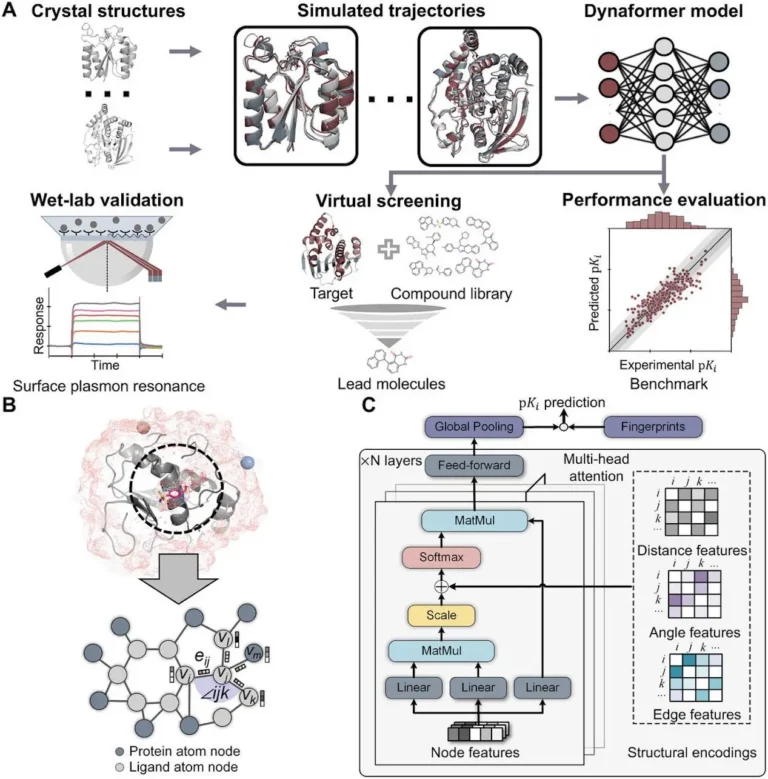
One major obstacle in structure-based drug design is accurately predicting the affinities of protein-ligand interaction. The accuracy of data-driven affinity prediction methods has improved recently, but its limitations remain. This is partly because these methods rely solely on static crystal structures, whereas thermodynamic ensembles between proteins and ligands typically dictate the real binding affinities. Using…
